
Researchers and Collaborators
Join our network of investigators and collaborators committed to improving health care for all.
High-quality and impactful research is built on a foundation of strong partnerships with regional leaders in health care innovation. Learn more about our process for establishing new studies, Virtua’s research policies, and how you can collaborate with us.
Please contact irb@virtua.org for applications, forms, templates, and guidance.
Research Process
Research Process (Getting Started)
Preparing Your Research Project
Before beginning your research project, it is important to complete several key steps to ensure compliance with institutional and regulatory requirements.
1. Determine Whether Your Project Meets the Definition of Research
Use our Research Determination Tool to assess whether your project qualifies as human subjects research and requires Institutional Review Board (IRB) review. Completing this step early can help you identify the appropriate regulatory pathway and avoid delays later in the process.
2. Complete Required CITI Training
All study personnel involved in the conduct of research must complete the required Collaborative Institutional Training Initiative (CITI) courses. Training requirements may vary depending on the type of research being conducted, the study population, and your role on the research team. Ensure your training is current before submitting an IRB application.
3. Assess Whether Additional Support Is Needed
If your project has funding, consider whether your project would benefit from support provided by the Research Administration and Operations (RAO) department. RAO can assist with study planning, regulatory requirements, budget development, contract review, operational feasibility assessments, and other research-related services.
4. Support for Principal Investigator (PI)-Initiated Studies
If you are conducting a PI-initiated study, early engagement with the appropriate research support teams is strongly recommended. These teams can provide guidance on protocol development, study implementation, regulatory documentation, and overall project management to help ensure a successful study launch.
5. Schedule an IRB Consultation
If you have questions about study design, IRB requirements, submission procedures, informed consent, or regulatory considerations, schedule a consultation with the IRB Office. An early consultation can help clarify requirements, address potential concerns, and streamline the review process.
Getting Started
Taking these preparatory steps before submitting your project will help ensure that your research is compliant, well-planned, and positioned for a timely review and approval process. Early planning and consultation can significantly improve the efficiency and success of your research study.
Virtua Research Flowchart
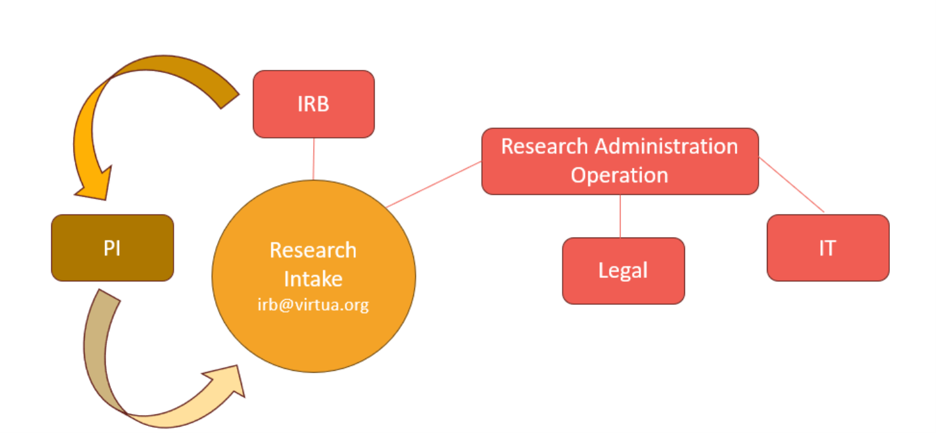
All research intake at Virtua Health, Inc. is done by Virtua Health IRB. Once a submission is received, the submission is screened for completion and the IRB will reach out directly to the study team for any additional questions or clarifications. In parallel, the research administration operation office will facilitate ancillary reviews including operational, legal, and IT requirements. After all requirements have been satisfied, the IRB may issue an approval letter to start your study. No official research may be initiated without IRB determination and/or approval.
IRB Review based upon Risk Level
The chart below will guide you to help assess the risk level of your research project.
| Full Board | Expedited | Exempt | QA/QI | Case Study/Non Human Research | |
|---|---|---|---|---|---|
| Risk | Greater than minimal risk to participants | Not greater than minimal risk to participants | Minimal risk | No added risk focused on improving internal processes or outcomes | Not applicable to human participants or does not involve human subjects as defined by regulations |
| Categories | Not covered under other review categories | Fits one of the 7 expedited review categories | Fits one of the 6 exempt categories | Activities designed for QA, QI, program evaluation, or operational improvement rather than generating generalizable knowledge | Activities involving non-human data/specimens or other projects that do not meet the regulatory definition of human subjects research |
| Examples | Interventions involving physical or emotional discomfort or collection of sensitive data | Biospecimen collection by noninvasive means; research with existing records/documents collected for non-research purposes in which participants are identifiable/deidentified (i.e., chart reviews) |
Research with totally deidentified record, anonymous surveys, no collection of PHI | Evaluating clinic workflow efficiency, assessing implementation of a new hospital process to improve patient care | Analysis of completely deidentified publicly available datasets, studies using non-human specimens, case study of a single patient, or laboratory projects that do not involve living individuals or identifiable private information |
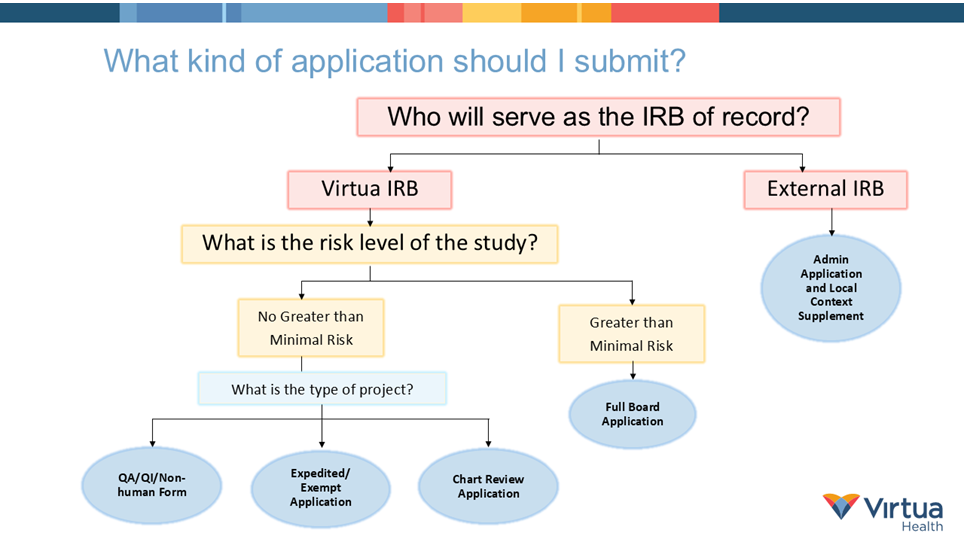
Application Type Decision Tree
The visual below may guide you to determine which application type to complete for your research project.
Conflict of Interest
One priority of our research is to manage conflict of interest for all involved parties. We provide the following resources to make a determination:
- Policies
- COI form for Investigators/ Co-Investigators
- COI form for study personnel
Application and Processing Fee Schedule:
| Review Type | Fee | Description |
|---|---|---|
| Full Board Review | $2,000.00 | Review of each new research protocol involving human subjects in externally funded studies. |
| Expedited Review | $1,500.00 | Covers initial review and preparation of regulatory documents, as well as ongoing reviews for amendments, adverse events, and annual reapprovals of externally funded studies |
| Local Context Review | $1,000.00 | Review for studies requiring local context review of externally funded studies. |
| Exempt Status Determination | $500.00 | Review conducted to determine qualification for exempt status of externally funded studies |
The IRB, at its discretion, shall designate some studies, e.g., cooperative group studies, single patient use, emergency use and internally funded protocols, as fee-exempt.
Additional Information
This application is intended only for studies that present Greater than Minimal Risk to participants. These are studies that involve interventions that can cause physical or emotional distress to the participant or collect sensitive data.
The following documents are required to complete the application.
- Signed Full Board Application
- Proposed informed consent
- Virtua HIPAA consent (if not included in the ICF)
- Relevant surveys, questionnaires, patients’ diaries, or data collection tools
- Investigator’s brochure, device manual or package insert, if applicable
- Participant recruitment procedures and materials, if applicable
For each investigator/co-investigator listed on the study application:
- Signed conflict of interest form
- Current curriculum vitae
- Verification of CITI Training
For Study Personnel
- Signed conflict of interest form
- Verification of CITI Training
*Vulnerable Populations
The vulnerable populations refers to but not limited to children, minors, pregnant women, fetuses, human in vitro fertilization, prisoners, employees, military persons and students in hierarchical organizations, terminally ill, comatose, physically and intellectually challenged individuals, institutionalized, elderly individuals, visual or hearing impaired, ethnic minorities, refugees, international research, economically and educationally disabled and healthy volunteers. If your study involves any vulnerable population, please download and submit Vulnerable Populations checklist with your IRB application.
This application is intended only for studies that either present no more than minimal risk to participants and fall into one of the expedited categories authorized by 46 CFR 46 and 21 CFR 56 or involve minimal risk to participants and fits one of the Exempt categories authorized by 45 CFR part 46, Subpart A.
The following documents are required to complete the application.
- Signed Expedited/Exempt application
- Paper Survey/Questionnaire Consent Template, if applicable
- Separate Data Collection Sheet, if applicable
For each investigator/co-investigator listed on the study application:
- Signed conflict of interest form
- Current curriculum vitae
- Verification of CITI Training
For Study Personnel
- Signed conflict of interest form
- Verification of CITI Training
*Vulnerable Populations
The vulnerable populations refers to but not limited to children, minors, pregnant women, fetuses, human in vitro fertilization, prisoners, employees, military persons and students in hierarchical organizations, terminally ill, comatose, physically and intellectually challenged individuals, institutionalized, elderly individuals, visual or hearing impaired, ethnic minorities, refugees, international research, economically and educationally disabled and healthy volunteers. If your study involves any vulnerable population, please download and submit Vulnerable Populations checklist with your IRB application.
This application is intended only for patient record reviews. By definition this will only apply to studies in which data is collected from patient’s medical records; either retrospectively or prospectively.
The following documents are required to complete the application.
- Signed Chart Review application
- Separate Data Collection Sheet, if applicable
For each investigator/co-investigator listed on the study application:
- Signed conflict of interest form
- Current curriculum vitae
- Verification of CITI Training
For Study Personnel
- Signed conflict of interest form
- Verification of CITI Training
*Vulnerable Populations
The vulnerable populations refers to but not limited to children, minors, pregnant women, fetuses, human in vitro fertilization, prisoners, employees, military persons and students in hierarchical organizations, terminally ill, comatose, physically and intellectually challenged individuals, institutionalized, elderly individuals, visual or hearing impaired, ethnic minorities, refugees, international research, economically and educationally disabled and healthy volunteers. If your study involves any vulnerable population, please download and submit Vulnerable Populations checklist with your IRB application.
This application is intended only for studies that are multisite studies and /or industry funded trials where Virtua Health IRB is relying on another IRB for ongoing review.
The following documents are required to complete the application.
- Signed Admin Review Application
- Approval from the IRB of record
- All study related document approved by the IRB of record
- Virtua HIPPA (if applicable)
For each investigator/co-investigator listed on the study application:
- Signed conflict of interest form
- Current curriculum vitae
- Verification of CITI Training
For Study Personnel
- Signed conflict of interest form
- Verification of CITI Training
*Vulnerable Populations
The vulnerable populations refers to but not limited to children, minors, pregnant women, fetuses, human in vitro fertilization, prisoners, employees, military persons and students in hierarchical organizations, terminally ill, comatose, physically and intellectually challenged individuals, institutionalized, elderly individuals, visual or hearing impaired, ethnic minorities, refugees, international research, economically and educationally disabled and healthy volunteers. If your study involves any vulnerable population, please download and submit Vulnerable Populations checklist with your IRB application.
In the event of an emergency use of an investigational drug, biologic or device on a human subject in a life-threatening situation in which no standard acceptable treatment is available, and in which there is not sufficient time to obtain IRB approval [21 CFR 56.102(d)]. The exemption, which may not be used unless all of the conditions described in 21 CFR 56.102(d) exist, allows for one emergency use of a test article without prospective IRB review. FDA regulations require that any subsequent use of the investigational product at the institution have prospective IRB review and approval.
DEFINITIONS:
Emergency use means the use of a test article on a human subject in a life-threatening situation in which no standard acceptable treatment is available, and in which there is not sufficient time to obtain IRB approval [21 CFR 56.102(d)].
https://www.ecfr.gov/current/title-21/chapter-I/subchapter-A/part-56/subpart-A/section-56.102
Test article means any drug for human use, biological product for human use, medical device for human use, human food additive, color additive, electronic product, or any other article subject to regulation under the act or under sections 351 or 354–360F of the Public Health Service Act [21 CFR 56.102 (i)].
https://www.ecfr.gov/current/title-21/chapter-I/subchapter-A/part-56/subpart-A/section-56.102
Life-threatening: for the purposes of section 56.102(d), includes the scope of both life-threatening and severely debilitating, as defined below.
- Life-threatening means diseases or conditions where the likelihood of death is high unless the course of the disease is interrupted and diseases or conditions with potentially fatal outcomes, where the end point of clinical trial analysis is survival. The criteria for life-threatening do not require the condition to be immediately life-threatening or to immediately result in death. Rather, the subjects must be in a life-threatening situation requiring intervention before review at a convened meeting of the IRB is feasible.
- Severely debilitating means diseases or conditions that cause major irreversible morbidity. Examples of severely debilitating conditions include blindness, loss of arm, leg, hand or foot, loss of hearing, paralysis or stroke.
When a potential participant lacks the capacity to provide informed consent, a legally authorized representative (LAR) may provide surrogate consent in accordance with federal regulations, New Jersey state law, the Access to Medical Research Act (N.J.S.A. 26:14 1 et seq.), and Virtua Health policy. Surrogate consent may only be used when the research cannot be feasibly conducted with individuals who can consent for themselves and when the IRB determines that the risk benefit profile is appropriate for this population.
Identification of a Legally Authorized Representative
Under the Access to Medical Research Act, an LAR is an individual legally permitted to make healthcare decisions for a person who lacks decision making capacity. Investigators must first assess and document the participant’s incapacity before seeking surrogate consent.
The following hierarchy, consistent with New Jersey law and the Access to Medical Research Act, must be used to identify an appropriate surrogate:
- Court appointed guardian
- Healthcare representative named in an advance directive
- Spouse, domestic partner, or civil union partner
- Adult child
- Parent
- Adult sibling
- Other adult relatives or individuals with a close personal relationship, as permitted under the Act
The investigator must document the surrogate’s authority and relationship to the participant.
Requirements for Surrogate Consent
The surrogate must receive the same information required for standard informed consent.
Under the Access to Medical Research Act, the surrogate must base decisions on:
- The participant’s known wishes, if previously expressed
- If unknown, the participant’s best interests
Investigators must document:
- The determination of incapacity
- The identity and legal authority of the surrogate
- The surrogate’s decision and signature on the consent form
Participant Assent
Consistent with the Access to Medical Research Act, investigators must seek the participant’s assent whenever possible. If the participant expresses resistance or dissent—verbally, physically, or behaviorally—the individual must not be enrolled or must be withdrawn unless the IRB has specifically approved an exception for compelling clinical or scientific reasons.
Ongoing Assessment of Capacity
Decision making capacity may change over time. Investigators must reassess capacity throughout the study. If a participant regains capacity, direct informed consent must be obtained as soon as feasible.
Post IRB Approval Requests
| Submission Type | When You Use It | Form |
|---|---|---|
| Continuing Report/Review | When a study is ongoing and requires annual IRB renewal to remain active. Please make sure to submit updated Signed conflict of interest form with the Continuing Review Report. |
Continuing/Closure Report |
| Closure Report | When the study is finished and no further IRB oversight is required. | Continuing/Closure Report |
| Progress Report | Providing progress updates for the study. | Continuing/Closure Report |
| Modification | When you need to change anything previously approved in the study e.g. protocol revisions, updates/addition to study documents, updates to study team members etc. | Study Modification Form |
| Unanticipated Adverse Events (UAEs) | When an event meets the criteria of UAE. | Reportable Event / SAE / UAE |
| Protocol deviations | A protocol deviation refers to any unplanned change, divergence, or departure from the approved study protocol. | Reportable Event / SAE / UAE |
| Protocol Violation | Protocol violation is a serious breach of the study protocol that can compromise participant safety, affect the reliability or completeness of study data, or violate regulatory requirements. | Reportable Event / SAE / UAE |
| Acknowledgment Request | Submitting a document that requires acknowledgment from the IRB. | Reportable Event / SAE / UAE |
| Reportable Events | AEs SAEs that meet the criteria of reporting or are required by the sponsor to be reported | Reportable Event / SAE / UAE |
| Data Safety Monitoring Report | Submitting Data Safety Monitoring Report to the IRB | Reportable Event / SAE / UAE |
| Marketing | All marketing related communications must follow Virtua guidelines | Marketing guidelines |
Please refer to Virtua policy to report to make sure that events are reported in a timely manner.
All forms must be signed by PI in order to be reviewed by the IRB.
If you do not have access to Virtua's Colleague Corner and need a document noted on this page, please contact us here.
Quality Assurance/Quality Improvement
For QA/QI project please submit the following
- Signed QA/QI assessment form
- Protocol
- Surveys/Data Collection Sheet, etc.
- If the PI or other Key Personnel is a medical student, resident or fellow, please upload letter from Graduate Medical Education Office confirming your dates of affiliation
- Please provide letter of support from the participating site documenting their knowledge and approval for this project
**All projects require submission to Virtua Health IRB for official determination as a QA, QI, or Case Report project.
Differences between Research and QA/QI
| Research | QA/QI | Non-Human / Case Reports | |
|---|---|---|---|
| Goal | To establish clinical practice standards where none are accepted OR to test a hypothesis. | To continuously assess or promptly improve a process, program, or system; OR improve performance as judged by accepted/established standards | To describe unique clinical observations, rare conditions, novel treatments, findings, or educational experiences; not intended to test a hypothesis or improve a system. |
| Initial Point | To answer a question or test a hypothesis | To improve already established standard | To document and share observations, experiences, or findings. |
| Benefits | Contributes to generalizable knowledge and may or may not benefit participants | Designed to promptly benefit a process, program, or system and may or may not benefit participants | Contributes to scientific or educational literature; generally, does not directly benefit subjects. |
| Risks/Burdens | May place participants at risk and stated as such. | By design, does not increase patient’s risk, with exception of possible privacy/confidentiality concerns | Minimal risk; may involve privacy/confidentiality concerns for case reports if identifiable information is included. |
| Data Collection | Systematic Collection of Data. | Systematic Collection of Data. | Uses existing clinical information, observations, records, specimens, or non-human data; not typically collected through a prospective systematic research protocol. |
| End Point | Answer a research question | Promptly improve a program/process/system | Describe and disseminate findings, observations, or educational information. |
| Testing/Analysis | Statistically prove or disprove a hypothesis | Compare a program/process/system to an established set of standards | Primarily descriptive; may include literature review or summary analysis but does not test a hypothesis. |
| Participation/Obligation | No obligation of individuals to participate | Responsibility to participate as component of care | No participant obligation |
| Effect on Program or Practice | Findings are generally not expected to immediately affect or change practice | Findings are expected to directly and immediately affect institutional practice | Findings may inform future research, education, or clinical awareness but are not intended to directly change institutional practice. |
Nursing Academic Research
Virtua supports academic and professional development. If your project involves a Nursing Academic degree, please contact AcademicAffiliations@virtua.org. More information can be found on the Academic Affiliation page of the Colleague Corner. Please log in to access this page.
Resources:
Helpful Links
- Belmont Report
- OHRP
- FDA: New Drug Development
- FDA: Medical Devices
- Human Subject Protection - 21 CFR Part 50
- Institutional Review Board - 21 CFR Part 56
- Protection of Human Subjects - 45 CFR Part 46
- FDA Information Sheet for PI and IRB
- OHRP Decision Charts
Policies
Templates
- Protocol Template (Interventional Research)
- Protocol Template (Non-interventional Research)
- Informed Consent Template
- Adverse Event Tracking Form Template
Tip Sheets
Need access to materials on this page?
If you do not have access to Virtua's Colleague Corner and need a document noted on this page, please contact us here.